DETDA Factory Production: Key Factors Behind High-Quality Curing Agents
2026-10-02
When it comes to high-performance epoxy curing, DETDA often steals the spotlight—but the real story starts long before it reaches the formulation. Behind every reliable batch lies a factory process where precision, quality control, and raw material integrity decide whether the curing agent truly delivers. Fortune, a name synonymous with production excellence, has turned these key factors into a science. Yet not all DETDA is created equal. So what separates a genuinely high-quality curing agent from the rest? The answer may surprise you—let's take a closer look at the production floor.
How Feedstock Sourcing Shapes the Final Color of Every Batch
The color of any batch begins long before the first drop is mixed—it starts in the fields, forests, or mines where raw materials are pulled from the earth. Two suppliers may sell what looks like the same pigment or resin, but subtle variations in soil composition, harvest timing, or extraction methods creep into the feedstock. These invisible fingerprints show up later as a warmer yellow, a duller red, or a haze that refuses to settle. Without strict sourcing protocols, even a trusted formula drifts batch to batch, leaving manufacturers chasing a shade they can't quite repeat.
What complicates matters is that feedstock isn't a static ingredient. Weather patterns shift the oil content in seeds, seasonal labor shortages slow harvests, and geopolitical friction reroutes supply chains overnight. A processor who suddenly switches from one regional supplier to another may think they're buying an equivalent product, but the molecular profile tells a different story. That difference cascades into oxidation rates, heat sensitivity, and ultimately, the final hue. Teams that treat sourcing as a fixed input rather than a living variable are constantly surprised by color drift.
The strongest operations treat feedstock mapping as part of the color development process itself. They run small-scale trials with incoming lots, document spectral data against origin, and build tolerance ranges that reflect real-world variability—not lab-perfect ideals. When a batch does shift, they can trace it back to a specific shipment, a field, even a week of rainfall. That level of traceability doesn't just stabilize color; it turns sourcing from a procurement headache into a competitive advantage that customers can see on the shelf.
The Nitration Step Where One Oversight Creates Off-Spec Isomers
In many fine chemical campaigns, the nitration stage is treated as a routine unit operation—until the isomer ratio drifts outside the accepted window. A single overlooked variable, such as a few degrees of extra exotherm during mixed-acid addition, can shift the ortho/para distribution enough to fail a downstream crystallisation spec. The result is not a failed reaction but a batch that quietly carries an off-spec impurity profile through to final isolation.
Operators often focus on total conversion while the real hazard hides in local mixing dynamics. When nitric acid is charged too rapidly into a sulfuric acid solution of the substrate, transient hot spots form before the agitator can homogenise the system. These pockets favour the thermodynamically more stable meta isomer or accelerate dinitration, even though the bulk temperature reading stays within range. The discrepancy only appears later, in HPLC traces that show a shoulder where the expected para peak should be clean.
Recovering from such a deviation is rarely straightforward. Once the off-spec isomer co-crystallises or co-distils with the desired product, simple reprocessing may not restore the required purity. The only reliable fix is to slow the addition profile, tighten acid strength tolerances, and install near-immediate reaction calorimetry—so the oversight that created the isomers is caught before it becomes a batch record entry.
Hydrogenation Timing and Catalyst Care Behind Low Residual Nitro
Residual nitro levels often trace back less to the hydrogenation step itself and more to how the catalyst was prepared and timed within the overall sequence. Rushing the reduction before upstream intermediates have fully converted or before the reactor atmosphere is properly inerted can leave stubborn nitro species that no amount of extended hold time will completely erase.
Catalyst care starts well before charging. Pre-wetting, avoiding exposure to air after activation, and matching agitation to particle size all influence whether surface sites stay active long enough to drive the reaction to completion. For sensitive substrates, adding catalyst in portions or adjusting temperature during the early stage can prevent localized over-reduction while still pushing residual nitro below the target limit.
Timing also matters in the work-up. Filtering too early traps partially reduced intermediates, while holding the reaction mixture too long after completion can degrade the product and mask the true endpoint. The usual practice is to sample at intervals once hydrogen uptake slows, then wait for two consecutive stable results before deciding the nitro has been fully consumed.
Vacuum Distillation Cuts That Separate Clear Product from Dimer Haze
The separation of clear product from dimer haze in vacuum distillation rarely follows a single sharp temperature reading. Under reduced pressure, the boiling gap between the desired monomeric cut and dimeric species can compress, and the haze boundary often shifts with slight changes in vacuum level or boil-up rate. Operators get better results by watching overhead vapor temperature alongside a timed takeoff profile, then pulling small samples across the suspected transition zone. A cut point set too late lets dimer-rich material slide into the main receiver and ruins clarity; a cut set too early sacrifices yield without meaningful quality gain.
A practical approach is to reserve an intermediate cut that begins when the first slight turbidity appears and ends before heavy haze develops. This transition fraction usually contains only a modest amount of the desired product but a disproportionate share of dimer and color bodies. It can be recycled into a later batch or redistilled under gentler conditions, keeping the main product clear. Because dimer formation is partially reversible at elevated temperatures, holding the pot at high temperature for an extended period will regenerate haze, so prompt cut switching matters more than achieving a perfectly dry endpoint.
In many towers, haze that appears mid-fraction is not caused by distillation curve overlap but by entrained liquid droplets carrying dimer-rich residue into the condenser. Demister pads, lower vapor velocity, and a short packed section above the feed point reduce this mechanical carryover. Combined with careful cut handling, these steps allow a clear product to be drawn over a wider range without the typical tail-end cloudiness. The result is a cleaner split between saleable clear material and dimer-laden heels, even when feedstock quality varies.
Keeping Moisture Out from Reactor Sampling to Drum Filling
Moisture rarely announces itself at the reactor stage—it slips in during a grab sample when a loose needle valve lets humid plant air backfill the headspace, or when a sample bottle sits open on the bench while the operator walks to the QC lab. Once that water finds a home in a moisture-sensitive intermediate, it tends to stay for the rest of the batch. The fix starts with closed-loop sampling systems that use septum-sealed ports and pre-dried vials, paired with a nitrogen sweep on the sample point so the line never sees ambient dew point. We also log the relative humidity at each sampling station and reject any sample taken when the local RH is above 35% unless the vessel is under positive nitrogen pressure.
Between the reactor and the drum, the biggest risk is condensation inside transfer piping that gets purged but not heated. A cold spot near a filter housing or a dead leg behind a block valve can collect enough moisture to fail a final KF spec, especially after a solvent swap or a long idle period. Replacing hard-piped sections with jacketed and traced lines, and adding dry-break couplings instead of open camlocks, keeps the product path sealed from the moment it leaves the reactor. We also push a slow nitrogen bleed through the transfer manifold for at least ten minutes before the first drum is connected, and we verify the line dew point with a handheld hygrometer rather than trusting a timed purge.
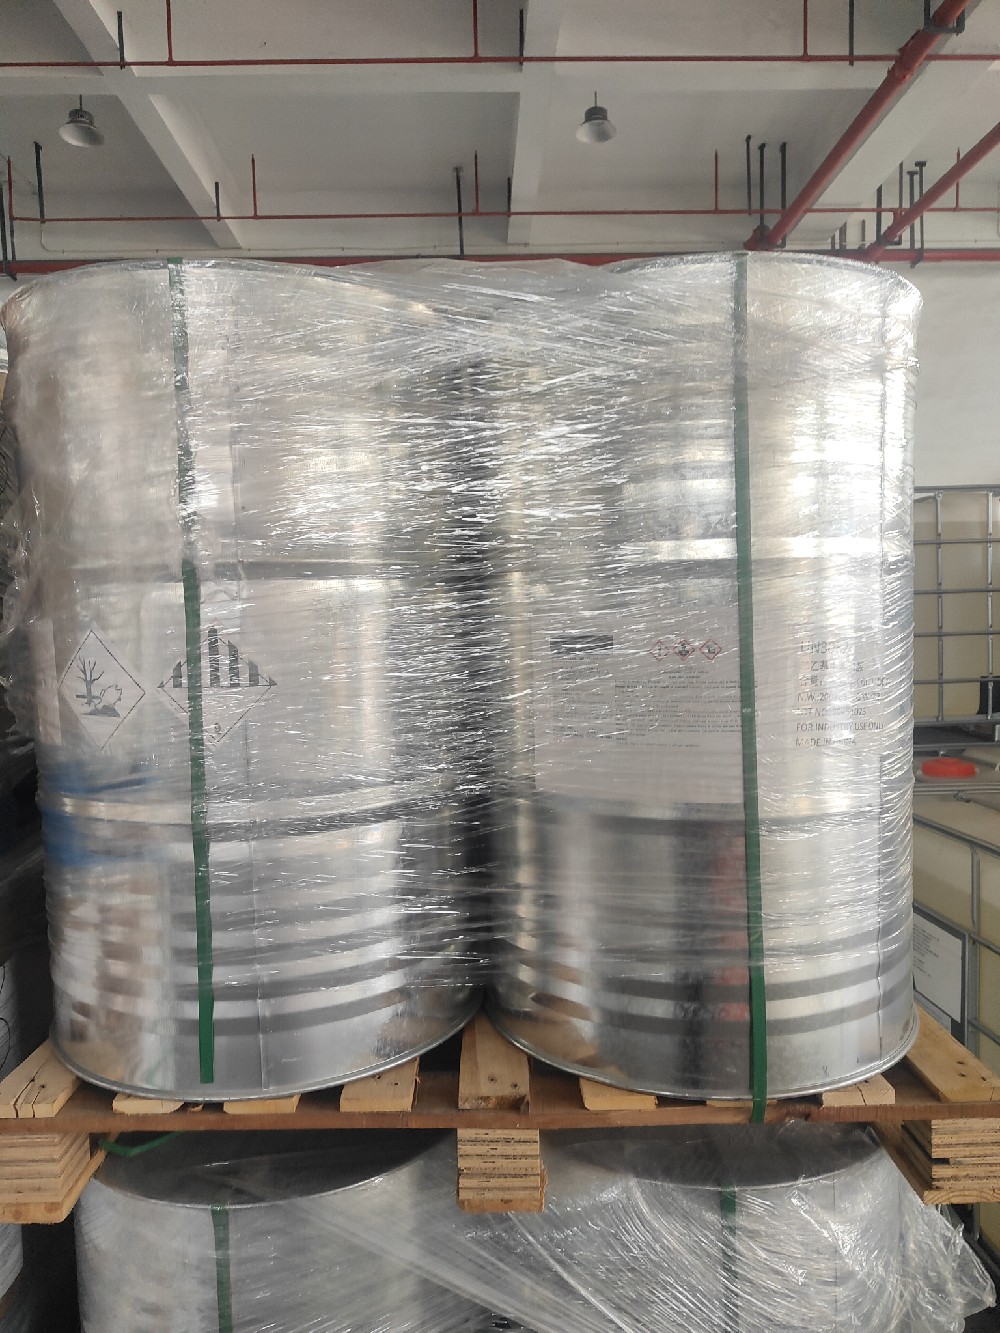
Drum filling is where all that earlier effort can be lost in a single unsealed lid. We fill through a dedicated stainless lance inserted into a 2-inch bung, with the displaced headspace vented through a desiccant-filled breather rather than an open vent. Each drum is pre-inerted with nitrogen to below 500 ppm oxygen and then checked for internal dew point before the fill starts. After filling, the closure is torqued under a nitrogen blanket, and a tamper-evident seal with an integrated moisture indicator goes on the bung. The result is a closed chain from reactor sample to sealed drum, with no point where ambient humidity gets a free pass.
Why Lab Tests Go Beyond Amine Value and Into Isomer Fingerprinting
Amine value has long served as a quick yardstick for cure behavior in epoxy systems, but it only tells you the total concentration of reactive amine groups. It says nothing about how those groups are arranged on the molecule. Two hardeners with identical amine values can behave very differently in a formulation because one might be mostly linear while the other carries bulky side chains or ring structures. Those structural differences shift gel time, exotherm profile, and even final crosslink density in ways a single titration number cannot capture.
Isomer fingerprinting picks up where amine value stops. By separating and identifying specific isomers—such as different methylene dianiline variants or positional isomers in cycloaliphatic amines—labs can flag batch-to-batch shifts that would otherwise go unnoticed. A supplier might change feedstock or distillation cuts, slightly altering the isomer ratio while keeping total amine content on spec. That hidden change can cause real problems downstream: longer induction periods, lower Tg, or unexpected viscosity drift. With fingerprint data, formulators catch those variations before they reach production, adjusting stoichiometry or rejecting the lot altogether.
The practical payoff shows up in high-stakes applications like adhesives for aerospace or electronics encapsulation. In those settings, a small shift in amine isomer distribution can mean the difference between passing thermal cycling and failing in the field. Relying on amine value alone is like checking the fuel gauge without looking at the octane rating—you know there is energy present, but not whether it will perform as expected. Isomer fingerprinting gives the molecular-level detail needed to keep performance consistent, making it an essential tool when total amine content is no longer enough to guarantee reliability.
FAQ
The purity of diethyltoluenediamine precursors and the control of isomer ratios directly influence color, viscosity, and reactivity. Even small amounts of moisture or unreacted intermediates can shorten shelf life and create inconsistent cure profiles.
Tight temperature ramps during the alkylation and reduction stages prevent side reactions that form dark, tarry byproducts. If the exotherm is not managed carefully, the amine value drops and the product tends to yellow faster in storage.
Distillation removes low-boiling impurities and residual catalysts that would otherwise interfere with epoxy crosslinking. Skipping or shortening this step often results in a wider cure window and lower mechanical strength in the cured resin.
Frequent sampling lets operators adjust hydrogen pressure, catalyst activity, and feed rates before deviations become irreversible. This is what separates a product that performs the same every time from one that drifts between lots.
Yes, and the effect is often underestimated. A selective hydrogenation catalyst reduces unwanted ring hydrogenation and keeps the product low-viscosity and light-colored, while a poorly chosen catalyst can raise odor and accelerate darkening.
Final filling is usually done under a dry nitrogen blanket in sealed lines. Moisture reacts with isocyanates and epoxies later, so even dew-point checks on the headspace are treated as a release criterion, not just a formality.
Beyond amine value and viscosity, gas chromatography tracks residual isomers, and color is measured against platinum-cobalt standards. Some producers also run a small-scale epoxy gel time test to confirm real-world curing behavior.
Absolutely. By adjusting the 2,4- and 2,6-isomer ratio, a factory can tune pot life and glass transition temperature for different epoxy systems, which is why the same base chemistry can serve both coatings and structural adhesives.
Conclusion
At the heart of every reliable DETDA batch lies a chain of decisions that start long before the reactor heats up. The choice of feedstock isn't just a procurement checkbox—it quietly dictates the final hue of the product, because trace impurities and aromatic substitution patterns carry through to the cured part. Then comes nitration, where a single lapse in temperature control or mixing can push the reaction toward unwanted isomers that later refuse to crystallize or cure uniformly. What separates a consistent factory from an average one is the discipline to treat these early steps not as routine chemistry, but as color and performance fingerprints already being written.
Downstream, hydrogenation timing and catalyst care decide how much residual nitro survives into the final drum. A tired catalyst or a rushed reduction leaves behind species that both darken the product and compromise pot life. Vacuum distillation then does the delicate work of separating clear, low-dimer DETDA from the hazy oligomers that form during storage. But even a perfect distillate can be ruined by moisture creeping in through sampling ports or drum-filling lines, so the best operations treat dryness as a production parameter, not a housekeeping note. Finally, lab testing has moved past simple amine value. Isomer fingerprinting—often by GC or HPLC with careful calibration—reveals the ratio of 2,4- to 2,6-isomers that actually governs reactivity and final mechanical properties. A high-quality curing agent isn't an accident; it's the sum of these interlocking controls, each one catching what the previous step left behind.
Contact Us
Contact Person: WANG SIHAI
Email: [email protected]
Tel/WhatsApp: +86-13815295628
Website: https://www.fortunechemtech.com/index_en.html